Chapter 42Analyzing Protein Structures Using an Ensemble Representation
This chapter discusses the mathematical basis and consequences of comparing an ensemble of unfolded-state molecules versus a given reference structure, and then presents the biological meaning of the structure comparison of unfolded (or intrinsically disordered) proteins at the ensemble level [1].
We address an important problem: Suppose an ensemble of structures is computed for a protein in an unfolded or disordered state (chapters 7 and 39), or even for a unimodal native state (chapters 15–18). Given a reference structure (for example, a crystal structure), or even multiple structures of exchanging states (chapter 27), how do we compare these reference structures to the ensemble? The answer will be surprising to many readers.
Mathematical Results
Terminology
Let and be the distance matrices of structures and , respectively. The distance root-mean-square deviation between distance matrices and is defined by
where denotes the Euclidean distance between atoms and in structure .
The Cartesian coordinate-based root-mean-square deviation between structures and is defined by
where denotes the Cartesian coordinates of the th atom in structure , and denotes the Euclidean -norm, that is, .
42.1.2 Results
Given an ensemble of structures , , and a reference structure , we have the following mathematical results:
where denotes the th structure in the ensemble , and denotes the distance matrix of structure (the same definitions for ), and stands for the ensemble average over all structures in the ensemble.
42.1.3 Brief Proof
Here we give an intuition of a proof sketch. See the Results section in Zagrovic and Pande [1] for a rigorous proof.
A metric space is a set with a distance function , such that for every ,
- with equality if and only if (nonnegativity);
- (symmetry);
- (triangle inequality).
By definition of dRMS, we can easily construct a metric space with distance function:
Now consider a simple case in this metric space, as shown in figure 42.1. Suppose we have an ensemble consisting of only two points, and , and a reference point . Let be the midpoint between and , then stands for the mean of the ensemble. Extend to such that . By triangle inequality, we obtain . Since and , we have . Since stands for the distance between the mean of the ensemble and the reference—that is, —and stands for the average over all distances between every point in the ensemble and the reference point—that is, —we thus have
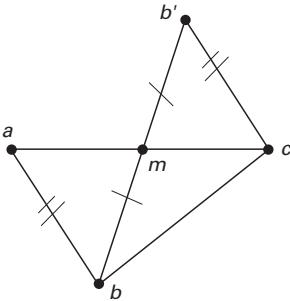
Figure 42.1 An illustration of the intuition. Credit: Michael Zeng and Bruce Donald.
It is easy to extend this intuition to a general case with ensemble size of more than 2.
Biological Significance
It has been observed that the mean unfolded-state distance matrices of an -helix are close to its corresponding native-state distance matrix. To exclude the effect of mathematical averaging, two tests of the -helical protein villin were performed in Zagrovic and Pande [1]:
Test 1 Use the members of the unfolded-state ensemble as reference structures instead of the native structure, and compare the (or RMSD) distance between the ensemble mean and each reference structure.
Results The native-state distance matrix is closer to the mean unfolded-state matrix than that of any member in the ensemble.
Test 2 Use native structures of other, unrelated proteins as reference structures, and perform a similar comparison.
Results The mean unfolded-state distance matrix is more similar to the corresponding native structure of the unfolded protein than any other native protein.
Reference
[1] B. Zagrovic and V. S. Pande. How does averaging affect protein structure comparison on the ensemble level? Biophysical Journal 87;4 (2004 Oct):2240–2246.